Sickle Cell Disease: A Comprehensive Clinical Guide to Pathophysiology, Diagnosis, and Management
Sickle cell disease (SCD) is a hereditary hemoglobinopathy caused by a point mutation in the β-globin gene, leading to the production of hemoglobin S (HbS). This comprehensive clinical guide covers the epidemiology, molecular pathophysiology, clinical manifestations, diagnostic approaches, and evidence-based management strategies for SCD, including acute crisis management, disease-modifying therapies, and curative options such as hematopoietic stem cell transplantation.
-
Introduction
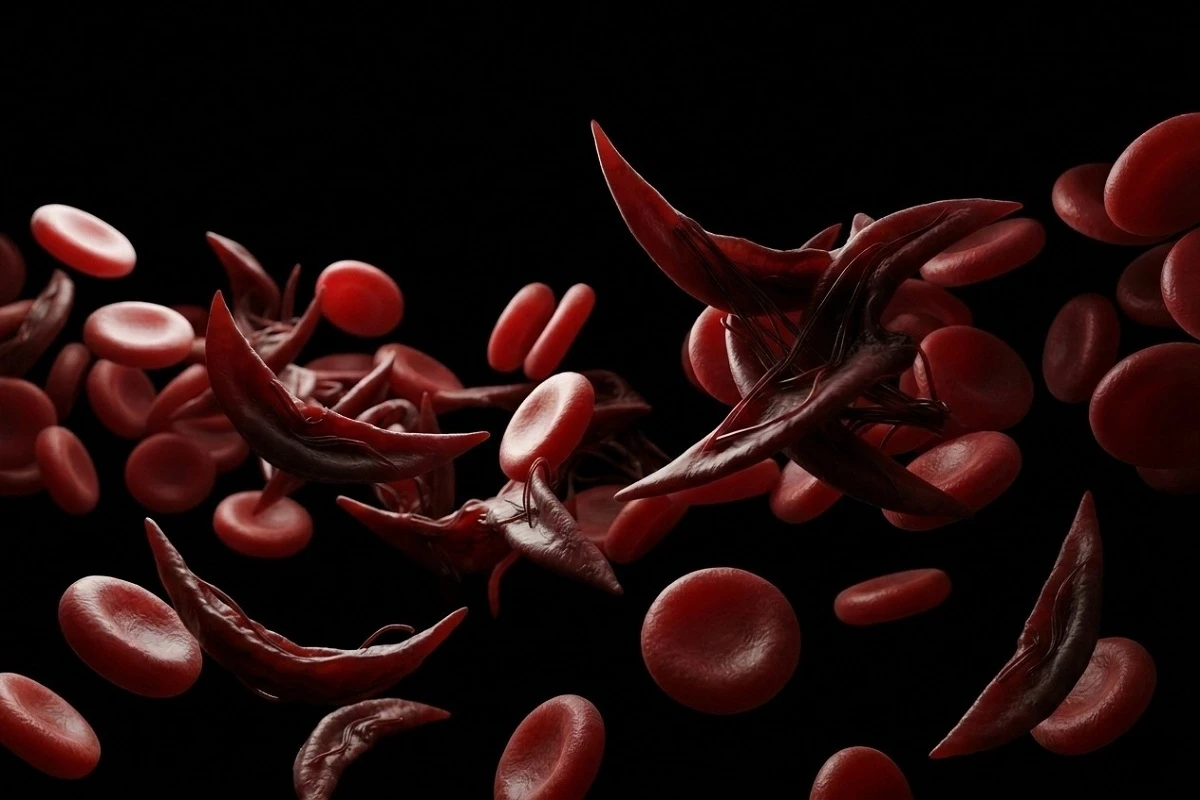
Sickle cell disease represents one of the most significant monogenic disorders worldwide, affecting millions of individuals across the globe. This inherited hemoglobinopathy is characterized by the production of abnormal hemoglobin S, which polymerizes under deoxygenated conditions, causing erythrocytes to adopt a characteristic sickle shape. These deformed cells lack elasticity, adhere to vascular endothelium, and occlude microcirculation, leading to recurrent episodes of pain, progressive organ damage, and reduced life expectancy.
The global burden of SCD is substantial, with an estimated 7.74 million affected individuals and approximately 515,000 new births annually. The disease predominantly affects individuals of African descent, though migration patterns have increased its prevalence in Western countries. Recent advances in understanding the pathophysiology of SCD, coupled with emerging therapeutic strategies including gene therapy, have transformed the clinical landscape for patients living with this challenging condition.
This comprehensive review provides clinicians with an evidence-based framework for understanding the molecular basis, clinical presentation, diagnostic approaches, and management strategies for sickle cell disease, with particular emphasis on acute complications and long-term preventive care.
-
Epidemiology and Global Burden
Prevalence and Distribution
Sickle cell disease affects populations worldwide, with the highest prevalence in sub-Saharan Africa, where carrier rates range from 10% to 40% in some regions. The HbS gene is carried by approximately 8% of the African American population, though individuals of any ethnicity can develop the disease. Other affected populations include those of Eastern Mediterranean, Middle Eastern, and Indian descent.
The global prevalence of SCD increased by approximately 41.7% between 2000 and 2021, with projections indicating continued growth, particularly in low- and middle-income countries. This increase reflects improved survival in childhood due to better preventive care, as well as population growth in regions where the disease is endemic.
Mortality Burden
The mortality burden of SCD is substantial and often
underestimated. In 2021, SCD caused 81,100 deaths in children under five years, making it the 12th leading cause of death in this age group when considering the total mortality burden. Traditional cause-specific mortality statistics significantly underestimate the true impact, with actual deaths estimated to be 11 times higher than reported figures. This disparity highlights the critical need for improved surveillance and healthcare delivery in affected regions.
Sickle Cell Trait
Sickle cell trait (HbAS) refers to the carrier state, where individuals inherit one sickle allele and one normal allele. Heterozygotes are typically asymptomatic carriers; however, complications can occur under extreme conditions such as high-intensity exercise, high altitude, or severe dehydration. Rarely, individuals with sickle cell trait may experience painless gross hematuria due to renal papillary necrosis, hyposthenuria, or exertional collapse and sudden death during extreme physical exertion.

Dr. Emmanuel Hirwa BAKUNDA is a Medical doctor and technology innovator dedicated to transforming medical education through digital solutions. With a clinical background in General pediatrics and child health, Dr. Bakunda combines frontline patient experience with a passion for health informatics and e-learning platforms.



Comments (0)